
MedDeviceGuide — IVD & Medical Device Knowledge Base
Medical Electrical Devices: Choosing an Operating Altitude Rating
Choose a defensible operating altitude for medical electrical equipment, with clearance calculations, thermal evidence, and separate EMS and aircraft assessments.
Medical Device SBOMs: Handling Missing Component Data From Suppliers
What to submit to FDA when a supplier won't or can't disclose SBOM component data: the sponsor's 524B duty, the justification route, purchasing controls, and evidence limits.
Changing a UK Responsible Person: Registration and Handover
How medical device manufacturers change a UK Responsible Person: MHRA registration steps, 2026 fees, designation contracts and the evidence to hand over in Great Britain.
Transport Damage: Connecting Complaint Photos to Packaging Validation
How RA/QA teams investigate medical device transport damage photos, satisfy QMSR and EU MDR complaint rules, and reconcile field damage against packaging validation.
Pilot Builds: Separating Engineering Learning From Saleable Product
Learn how to classify medical device pilot builds under FDA QMSR and ISO 13485, separate engineering evidence from saleable product, and handle investigational distribution.
Device Rework: Defining the Instruction and Reverification
Learn how to author compliant medical device rework instructions, determine reverification scope under ISO 13485 clause 8.3.4 and QMSR 820.3(b), and avoid undocumented fixes.
Reusable Device Cycle Limits: How to Justify Reprocessing Claims
Compare FDA and EU MDR routes for reusable device life: when to claim a fixed reprocessing cycle count versus end-of-life criteria, and what aging evidence regulators accept.
Medical Device Kits: Who Assembles, Labels, and Takes Responsibility?
A regulatory map assigning FDA establishment registration, 21 CFR 801.1 labeling, UDI labeler status, 510(k) discretion, and QMSR duties across kit assemblers and specification developers.
Brazil Device Registration Transfers: What Must Move With the Holder?
Under Brazil RDC 903/2024, only Class III/IV registrations transfer while Class I/II notifications restart. Here are the paired filings, 180-day clock, and labeling rules.
Device Label Durability: Testing the Actual Cleaning Exposure
How to build an audit-ready medical device label durability evidence chain against validated cleaning and disinfection cycles rather than generic peel tests.
IVD Specimen Stability: Matching the Claim to Collection and Transport
Support IVD specimen stability claims under 21 CFR 809.10 and EU IVDR Annex II by aligning labeled collection, additives, hold times, and transport with verified study configurations.
When Device Software Support Ends: The Customer Transition Plan
How medical device manufacturers structure customer transition plans when software reaches end of support, mapping required evidence, IMDRF milestones, and surviving duties.
Can a Discontinued Device Still Be a 510(k) Predicate?
FDA permits substantial equivalence claims to devices no longer marketed in the U.S. Here is how to verify eligibility, satisfy 21 CFR 807.87(f), and build evidence without physical samples.
One Test Report, Several Device Models: Proving Variant Coverage
How FDA guidance and the EU MDR decide which device models one test report covers, and which records justify an untested size or configuration.
IVD Reference Intervals: Established Values vs Local Verification
When a laboratory may adopt an IVD manufacturer's reference interval, when CLIA requires verification or establishment, and what ISO 15189 adds only for accredited laboratories.
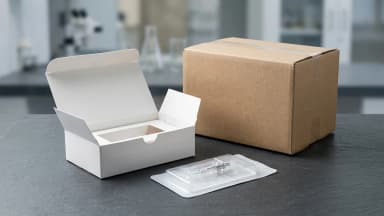
Which Device Packaging Levels Need Their Own UDI Carrier?
A practical decision guide to UDI placement across medical device packaging levels under FDA, EU MDR, and TGA rules, covering carriers, unique identifiers, and shipper exemptions.
Canadian Medical Device Licence Changes: Which Filing Path Applies?
How to classify Canadian medical device licence modifications into significant changes, statutory amendments, minor REP transactions, or annual renewals under SOR/98-282.
AccessGUDID vs FDA Listing: Which Record Answers Your Question?
Compare AccessGUDID and FDA's establishment registration and device listing database to choose the correct public record for device identifiers, establishments, and listing status.
Is This Combination an MDR Device or a Medicinal Product?
A manufacturer guide to EU MDR Article 1(8) and 1(9): classify integral ancillary devices, medicinal products, and administration devices with the four-scenario identity worksheet.
Does This Change Need a New 510(k), or Does a PCCP Cover It?
Mark whether one 510(k) device change still owes a new filing, is covered by a cleared 515C PCCP, or is documented without a 510(k).